Unveiling the Role of FOXF1 Variants in Pulmonary Arterial Hypertension Pathogenesis
Highlight
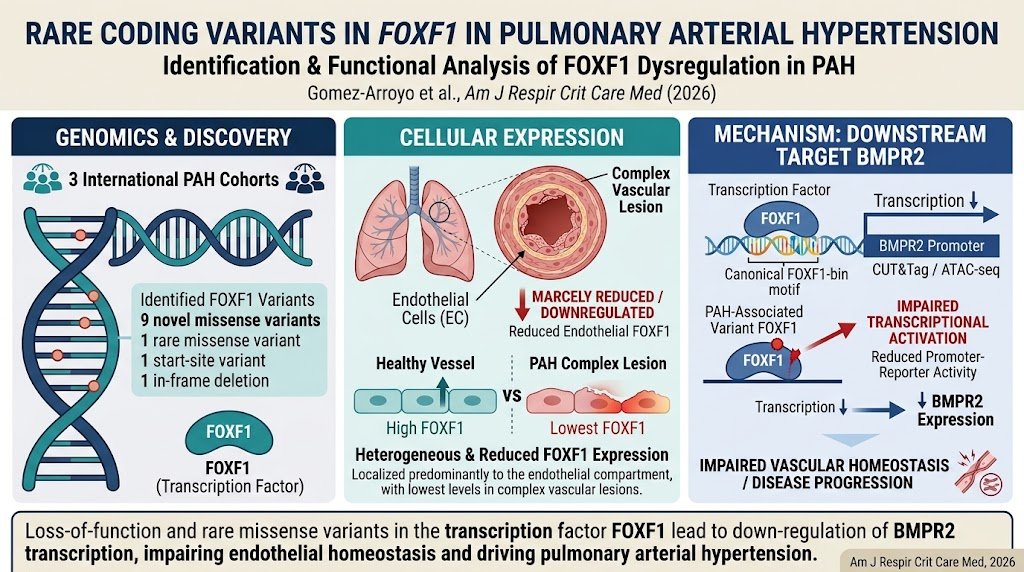
- Identification of ultra-rare missense and coding variants in FOXF1 among PAH patients from international cohorts.
- FOXF1 expression is predominantly endothelial and significantly reduced in PAH lungs, particularly in abnormal vascular lesions.
- Functional studies demonstrate FOXF1 directly regulates BMPR2 transcription, a known PAH-associated gene.
- Dysfunction of FOXF1 variants results in diminished transcriptional activity, suggesting a pathogenic mechanism for PAH.
Study Background
Pulmonary arterial hypertension (PAH) is a progressive vasculopathy characterized by elevated pulmonary arterial pressure leading to right heart failure and premature mortality. While heritable forms of PAH are linked to pathogenic variants in genes such as BMPR2, the majority of nonheritable PAH cases lack identifiable genetic causes. Emerging data suggest that beyond large-effect loss-of-function mutations, rare missense coding variants might contribute to disease risk by subtly impairing protein function or regulation. FOXF1 encodes a conserved transcription factor critical for lung vascular development and endothelial cell function. Elucidating the contribution of FOXF1 variants could improve genetic diagnosis and mechanistic understanding of PAH.
Study Design
The investigation included three independent, international cohorts of PAH patients screened for rare, predicted deleterious variants in the FOXF1 gene. Genetic data analysis focused on ultra-rare missense substitutions and in-frame deletions. FOXF1 expression was assessed in human lung tissue using bulk and single-cell RNA sequencing, supplemented by immunofluorescence. Bioinformatics integrated chromatin profiling (CUT&Tag, ATAC-seq) to identify FOXF1 binding motifs and downstream gene targets. Functional validation was performed via promoter-reporter assays in vitro comparing wild-type FOXF1 to patient-derived variants.
Key Findings
Analysis across cohorts uncovered ten missense variants, one start-site variant, and one in-frame deletion in FOXF1 in PAH patients but not in controls. Transcriptomic and histological studies revealed reduced FOXF1 expression predominantly in lung endothelial cells of PAH patients, with marked suppression in endothelial cells located within complex vascular lesions. This pattern was distinct from other chronic lung diseases, underscoring specificity for PAH.
Chromatin assays mapped a canonical FOXF1-binding sequence within the promoter region of BMPR2, a gene with established pathogenic variants in hereditary PAH. Promoter-reporter assays demonstrated that wild-type FOXF1 enhances BMPR2 transcription, whereas patient-associated FOXF1 variants impaired this transcriptional activation.
Together, these findings indicate that FOXF1 variants likely contribute to PAH pathogenesis by disrupting the transcriptional regulation of BMPR2 and possibly other vascular genes, leading to dysfunctional endothelial signaling and vascular remodeling.
Expert Commentary
This study advances our understanding of PAH genetics by implicating FOXF1 as a novel susceptibility gene and offering mechanistic insight through BMPR2 regulation. The specific downregulation of FOXF1 in PAH endothelial cells within lesions suggests a pathobiological role confined to diseased vasculature. While findings are compelling, limitations include relatively small variant counts and lack of in vivo functional models. Further studies to characterize how FOXF1 variants alter endothelial cell biology and to explore therapeutic targeting of FOXF1-driven pathways will be invaluable.
Conclusion
This comprehensive genomic and transcriptomic analysis identifies rare FOXF1 coding variants as contributors to PAH susceptibility. The data implicate FOXF1 dysregulation and its downstream effect on BMPR2 transcription as critical components of disease pathogenesis in a subset of PAH patients. These insights enrich the genetic landscape of PAH and suggest new avenues for molecular diagnostics and targeted intervention.
Funding and ClinicalTrials.gov
The study was supported by international collaborative grants and institutional funding from contributing centers. No specific clinical trial registration applies.
References
1. Gomez-Arroyo J, et al. Rare Coding Variants in FOXF1 are Associated with Pulmonary Arterial Hypertension. Am J Respir Crit Care Med. 2026 Oct 10; PMID: 42858161.
2. Machado RD, et al. Genetics and genomics of pulmonary arterial hypertension. Genome Med. 2015;7(1):99.
3. Morrell NW, et al. Targeting BMP signalling in pulmonary arterial hypertension. Eur Respir J. 2016;47(1):1-13.
4. Rajasekaran S et al. FOXF1 expression and function during lung vascular development. Vasc Cell. 2017;9:9.
This article was created using several editorial tools, including AI, as part of the process. Human editors reviewed this content before publication.